EducationHow a peptide is made
How Peptides Are Made: Solid-Phase Synthesis Explained

Written by Dr. Sufia Ul Haque
Chemist and scientific writer · Published · 5 min read
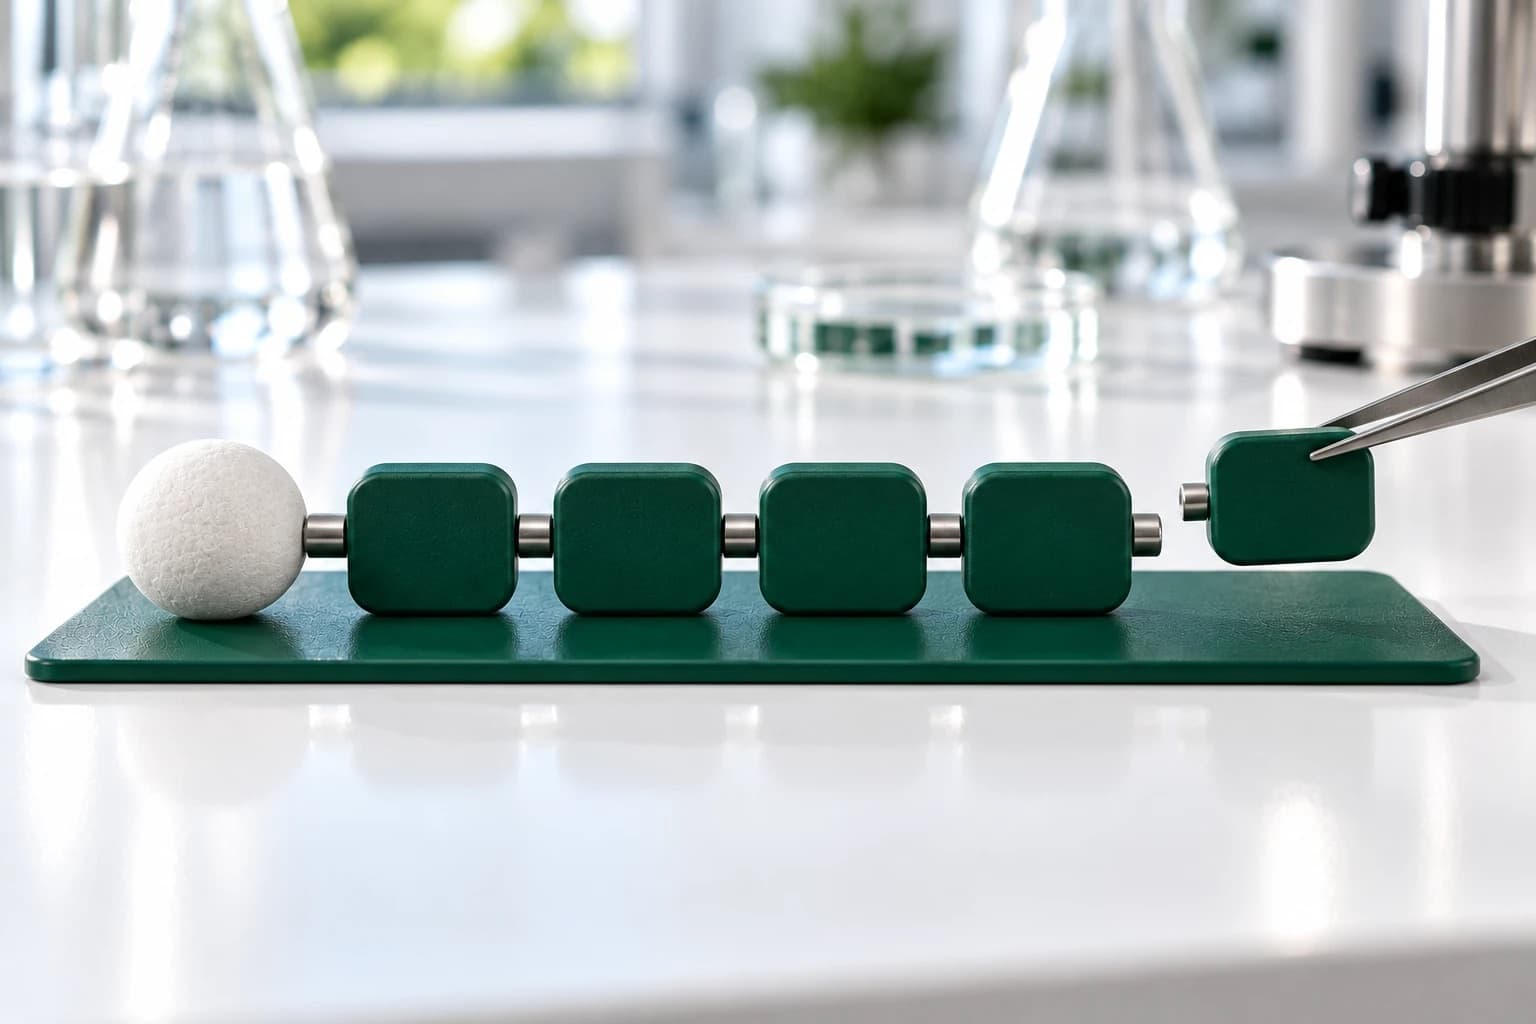
Solid-phase peptide synthesis (SPPS) builds a peptide one amino acid at a time while the growing chain remains attached to an insoluble resin. The unexpected part is its direction: conventional SPPS starts at the C-terminus and works toward the N-terminus, opposite to the way peptide sequences are normally written.
In this article+−
Where did solid-phase peptide synthesis begin?
In 1963, chemist Bruce Merrifield reported a method for assembling a tetrapeptide, Leu-Ala-Gly-Val, while it was attached to a polymer support. He later received the 1984 Nobel Prize in Chemistry for developing chemical synthesis on a solid matrix. His key insight was practical: if the growing molecule stays attached to an insoluble material, excess reagents and reaction by-products can be washed away without isolating the peptide after every step.
That idea transformed a demanding sequence of separate reactions into a repeatable process. Modern synthesis uses different protecting groups and cleavage chemistry from Merrifield’s original demonstration, as Ghosh and Lubell explain in their review. The central principle remains the same: keep the chain attached, perform the next reaction, and remove what is no longer needed.
My background in organic synthesis makes this approach especially interesting. In multistep synthesis, purification between reactions can take considerable time. Solid support does not eliminate the need to control chemistry, but it changes how intermediate material is handled.
Why is a peptide assembled from C to N?
A peptide sequence is conventionally written with its amino, or N-terminal, end on the left and its carboxyl, or C-terminal, end on the right. Conventional SPPS begins with the C-terminal amino acid attached to a resin through its carboxyl group. Its amino group can then react with the next incoming amino acid. As the chain lengthens, each new residue is added toward the N-terminus.
RelatedWhich end of a sequence is which
The incoming amino acid cannot simply react at every available site. Its amino group is temporarily protected, while its carboxyl group is activated for coupling. Reactive side chains may also need protecting groups so they do not form unwanted bonds. After each addition, the temporary N-terminal protection is removed to expose the site for the next residue.
This C-to-N arrangement is the established approach, not a universal chemical rule. Researchers have also developed N-to-C synthesis methods, although these remain less common, as Ghosh and Lubell discuss.
What happens during each synthesis cycle?
A typical cycle has three main parts. First, deprotection removes the temporary protecting group from the growing chain’s amino end. Next, coupling joins a suitably activated, protected amino acid to that free end. Finally, washing removes excess reagents and soluble by-products. The cycle repeats until the planned sequence is assembled.
The chain stays attached to the resin
One cycle, three parts
- Deprotect. Remove the temporary protecting group from the growing chain's amino end.
- Couple. Join a suitably activated, protected amino acid to that free end.
- Wash. Remove excess reagents and soluble by-products.
Repeat until the planned sequence is assembled.
Two common strategies are called Fmoc and Boc, named for their temporary amino-protecting groups. Fmoc protection is generally removed with a base such as piperidine; acid-labile side-chain groups are usually removed later. Boc protection is removed with acid, commonly trifluoroacetic acid (TFA), while final cleavage in classical Boc methods can require stronger acidic conditions. Behrendt and colleagues describe Fmoc as the dominant modern approach, with Boc retained for selected applications.
Actual protocols may add coupling tests, repeated couplings or capping steps. Capping can block unreacted amino groups from participating in later couplings, limiting further extension of those chains. These checks and controls matter because a reaction that looks complete on paper may leave some chains behind.
How is the completed peptide released?
After the last amino acid has been added, the peptide must be separated from the resin and, where necessary, freed from its side-chain protecting groups. In many Fmoc-based methods, a TFA-containing cleavage mixture performs both jobs. Scavengers in that mixture help capture reactive fragments released during deprotection, limiting unwanted side reactions.
TFA may also remain associated with basic groups on the peptide as a negatively charged counter-ion. Sikora and colleagues explain why peptide preparations can therefore contain trifluoroacetate and why counter-ion exchange may be used to obtain another form, such as acetate. Not every peptide has the same counter-ion composition, and retained TFA does not always represent a simple fixed-ratio salt.
RelatedWhy the salt form changes the molecular weight
For someone reading a peptide specification, this distinction matters. The amino acid sequence tells you what chain was assembled, but the isolated material may also contain counter-ions, water or other components that affect its measured mass.
Why do small reaction losses become important?
Each coupling has a chance of being incomplete. If a chain misses a residue but continues to grow, the result is a deletion sequence. If growth stops, the material may contain a truncated peptide. Other side reactions can produce modified sequences. Merrifield described these broad impurity categories in his Nobel lecture.
Consider a deliberately simplified calculation: if every coupling succeeds with 99% efficiency, the fraction of full-length chains after 14 couplings is 0.99¹⁴, or about 87%. After 49 couplings, it is about 61%. These figures are illustrations, not measured yields or predictions for a real peptide. Actual efficiency depends on sequence, chemistry, monitoring and repeated coupling steps.
A deliberately simplified calculation
Take a coupling that succeeds 99 times in 100, and assume nothing else goes wrong.
After 14 couplingsabout 87%
of chains are full length, about 87 percent
After 49 couplingsabout 61%
of chains are full length, about 61 percent
Arithmetic, not a measured yield
In my experience with organic synthesis and chromatography, a clean-looking reaction scheme does not guarantee a single product in the final sample. Analytical characterization is essential because closely related compounds can coexist even when the intended transformation is well understood.
RelatedWhat the impurities in a purity figure are
How is a crude peptide purified and checked?
Once the peptide is released, the crude mixture may contain the desired sequence alongside deletion, truncated or chemically modified products. Preparative reversed-phase high-performance liquid chromatography (HPLC) is a widely used purification method. It separates compounds according to how they interact with a stationary phase and a moving solvent. Ion-exchange methods can also be useful.
Some impurities are difficult to separate because their chemical properties closely resemble those of the target peptide. That is why chromatographic purification and analytical confirmation serve different purposes: one enriches the desired material, while the other helps establish what is present. The purified peptide is often isolated by lyophilization, or freeze-drying.
RelatedHow freeze-drying works
Long sequences may be assembled from shorter peptide segments using native chemical ligation, rather than relying on one uninterrupted synthesis. Recombinant expression offers another route for suitable sequences. The choice depends on the peptide and the chemistry needed.
Frequently Asked Questions
What is the difference between Fmoc and Boc synthesis?+−
Both use temporary amino-protecting groups, but they are removed under different conditions. Fmoc is usually removed with base, whereas Boc is removed with acid. The final cleavage conditions also differ. Fmoc is widely used in contemporary SPPS; classical Boc methods remain useful for certain specialized sequences and applications.
Why do synthetic peptides sometimes contain TFA?+−
Trifluoroacetic acid is often used during cleavage or purification. Peptides containing basic groups may retain trifluoroacetate as a counter-ion, and additional TFA can sometimes remain in a preparation. The amount depends on the material and process. Counter-ion exchange may replace it with acetate or another suitable ion.
Can solid-phase synthesis make peptides of any length?+−
There is no single fixed length limit. However, incomplete reactions and side reactions can accumulate as a chain grows, making longer sequences harder to obtain cleanly. For some targets, researchers synthesize shorter segments and join them by ligation. The best route depends on the sequence and required modifications.
Sources
- Merrifield RB. Solid phase peptide synthesis. I. J Am Chem Soc (1963). doi.org/10.1021/ja00897a025
- Nobel Prize in Chemistry 1984; Merrifield Nobel lecture. nobelprize.org/prizes/chemistry/1984/summary
- Ghosh K, Lubell WD. N- to C-peptide synthesis. J Pept Sci (2025). doi.org/10.1002/psc.70019
- Mäde V et al. Automated solid-phase peptide synthesis. Beilstein J Org Chem (2014). doi.org/10.3762/bjoc.10.118
- Behrendt R et al. Advances in Fmoc SPPS. J Pept Sci (2016). doi.org/10.1002/psc.2836
- Paravizzini SJ et al. Backbone protecting groups. Angew Chem Int Ed (2025). doi.org/10.1002/anie.202509939
- Kekessie I et al. Process mass intensity in peptide manufacturing. J Org Chem (2024). doi.org/10.1021/acs.joc.3c01494
- Lamb CMG, Shi J, Wilden JD, Macmillan D. Novel electrochemically-mediated peptide dethiylation in processes relevant to native chemical ligation. Org Biomol Chem. 2022;20:7343–7350. doi.org/10.1039/D2OB01499H
- Sikora K et al. The role of counter-ions in peptides. Pharmaceuticals (2020). doi.org/10.3390/ph13120442
- Cistrone PA, Bird MJ, Flood DT, et al. Native chemical ligation of peptides and proteins. Curr Protoc Chem Biol. 2019;11(1):e61. doi.org/10.1002/cpch.61
About the author
Dr. Sufia Ul Haque
Chemist and scientific writer · PhD in chemistry, Aligarh Muslim University
Dr. Sufia Ul Haque is a chemist and scientific writer. She holds a PhD in chemistry from Aligarh Muslim University and carried out postdoctoral research in organic and natural-product synthesis in the United States, where her work included multistep synthesis, purification and chromatography. She has published in journals including Biosensors and Bioelectronics, the Journal of Power Sources and Scientific Reports, and reviews for several international journals. She writes about chemistry for readers without a scientific background.
Disclosure: No conflicts of interest declared.
The author checked the claims in this article against its sources and approved the text as published on .